Skip to content
  • 4k display support?

    SAMSON
    2
    1
    1 Votes
    2 Posts
    2k Views
    StephaneS
    Hello, high-resolution displays are not supported natively yet, but Windows 10 indeed allows you to change the settings. On my 4K monitor, I use the method explained there : https://blog.pauby.com/post/high-dpi-settings-windows-10/ , but I choose "System" instead of "Application". My system DPI adjustment is at 200% in Windows, and here is how it looks like: [image: 0_1612374085845_ce0855be-0ca8-49e4-a8fd-bbf45f42f1b6-image.png] Could you please let me know if this works for you?
  • GROMACS files not seen

    SAMSON Extensions
    7
    0 Votes
    7 Posts
    4k Views
    S
    there is RMSF and hydrogen bond
  • [User Support] Adenita

    SAMSON Extensions
    2
    2 Votes
    2 Posts
    2k Views
    O
    I have a structure I would like to alter a bit using Adenita, but whenever I try to select it, it becomes non-responsive and crashes. I've tried using a .json file and a .pdb file and neither is capable of having parts selected. I tried on two different computers, one with a quad-core and one with a 6-core and it still crashed. Is there a way I can perform this on my build? Or is this a software issue that others experienced? OS: Windows 10 Ver: SAMSON 0.8.5
  • 0 Votes
    2 Posts
    2k Views
    DmitriyMarinD
    Dear @Olivia , The Adenita extension has been developed and is supported by external developers ( @Elisa ). And Adenita is in an alpha stage and might have some issues. We plan on porting this extension to the most recent SAMSON version. You can also copy your question to the specific topic: Adenita - User Support.
  • Adenita not showing up

    SAMSON Extensions
    4
    0 Votes
    4 Posts
    2k Views
    DmitriyMarinD
    Great! Thank you for your reply!
  • Autodock Constraints

    SAMSON Extensions
    4
    0 Votes
    4 Posts
    3k Views
    StephaneS
    Dear Sebastian, one possibility might be to ask the Vina extension to compute many modes (e.g. 1000) and then use a script to filter out the conformations that do not satisfy the distance constraint. Another possibility might be to directly modify the source code of Vina to add a term to the scoring function that would penalize one or more unsatisfied constraint(s) (similar to a NMR energy function). We're going to look into this. Thanks for the suggestion. Best, Stephane
  • Autodock Vina Extended problem

    SAMSON Extensions
    23
    0 Votes
    23 Posts
    26k Views
    S
    Thanks this is very good to know
  • Background/Visualization issues on 2015 Macintosh

    SAMSON
    3
    1 Votes
    3 Posts
    2k Views
    AdminA
    Hi Nick, thanks for the posts, we've sent you messages about how to reach us for chat so we can help you.
  • Ligand Receptor Interaction

    SAMSON Extensions
    11
    0 Votes
    11 Posts
    7k Views
    DmitriyMarinD
    Dear @Mariangela , To find and show hydrogen bonds you can use the Hydrogen Bond Finder Extension. It allows you to find all H-bonds in the selected system or between two systems, for example, between a receptor and a ligand. To find/show H-bonds between a receptor and a ligand, perform the following steps (see the gif at the end): in the Hydrogen Bond Finder Extension click on "...in the current selection and the system"; select the receptor in the Document view and in the Hydrogen Bond Finder click Set; [image: 0_1607705913844_07ee1a78-6dba-4dcc-912b-bf40a6d8a6fa-image.png] set the Settings of the Hydrogen Bond Finder if necessary; now, select only the ligand in the Document view and in the Hydrogen Bond Finder click Add hydrogen bond visual model Now you should have a new visual model in the Document view. If you select it, you can modify its options in the Inspector. See an example below: [image: alt text]
  • Energy profile from P-NEB

    SAMSON Extensions
    3
    0 Votes
    3 Posts
    2k Views
    V
    Yes, it would be useful to have the energy profile along the path. Not only the height of the barrier, but also its width, shape, etc.
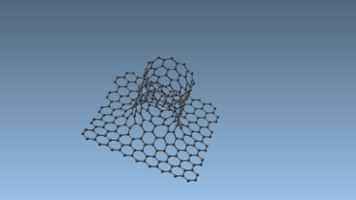
  • How to do a carbon nanotube attached in a graphene sheet?

    Modeling
    3
    2
    0 Votes
    3 Posts
    4k Views
    DmitriyMarinD
    Alternatively, you can do this without the Brenner interaction model but with the built-in minimization which uses the Universal Force Field (UFF) and the Add editor (shortcut: A) (it's Carbon by default). First, you will need to modify SAMSON preferences (Ctrl+K): -1- For the Add editor, disable the "Adjust hydrogens ..." option. [image: 0_1600418725406_Preferences-Add-no-adjust-H.png] -2- For the Minimization, disable the "Warn me when hydrogens appear to be missing" warning to prevent it from poping-up on each new bond connection. [image: 0_1600418739623_Preferences-Minimize-no-H-warning.png] Then you can apply the minimization: Home >Simulate > Minimize. Start connecting atoms in the nanotube and the graphene sheet using the Add editor (A) - to create a bond simply connect two atoms with this editor.
  • Increase font and icons size

    Getting started
    6
    1
    0 Votes
    6 Posts
    5k Views
    OneAngstromO
    @aaa Thank you for reporting your solution here!
  • GROMACS Wizard doesn't show

    SAMSON
    2
    0 Votes
    2 Posts
    2k Views
    AdminA
    Hi James, thanks for reporting this. We sent you an email to determine the issue.
  • Problems with the visualization of molecules

    SAMSON
    5
    1
    0 Votes
    5 Posts
    4k Views
    M
    Okay, Thanks.
  • Gromacs Wizard Membrane Protein

    SAMSON Extensions
    4
    0 Votes
    4 Posts
    3k Views
    AdminA
    The error message ("atom N not found in buiding block 1ACE while combining tdb and rtp") suggests that, in your model, 1ACE contains a N that should not be there (according to GROMACS topologies). You might have to remove it, or move it to the next residue. You can reorganize atoms, residues, etc. easilyby just dragging nodes around in the Document View.
  • samson core exe is not runing in windows 7

    Getting started
    2
    0 Votes
    2 Posts
    2k Views
    StephaneS
    Hello, could you please tell us which graphics card you have? This seems to indicate a GPU issue.
  • 0 Votes
    3 Posts
    2k Views
    StephaneS
    Indeed in the current version of the Gromacs Wizard, you will need to have a force field which allows for both biomolecules and CNTs. Then you can add such a force field in the settings of the module. However using a 'one-shot parameterization' (e.g. using the ATB server) will not work at the moment with the Gromacs Wizard.
  • Can't load CIF files

    SAMSON Extensions
    10
    0 Votes
    10 Posts
    6k Views
    M
    @dmitriymarin @admin It actually was this. Thanks for solving the problem!
  • Setting pivot point in Arap path

    SAMSON Extensions
    5
    0 Votes
    5 Posts
    3k Views
    E
    Thank you, Stephan. I will give it a try.
  • Docking results export

    SAMSON Extensions
    17
    0 Votes
    17 Posts
    11k Views
    H
    Dear @dmitriymarin thank you very much. It works.